
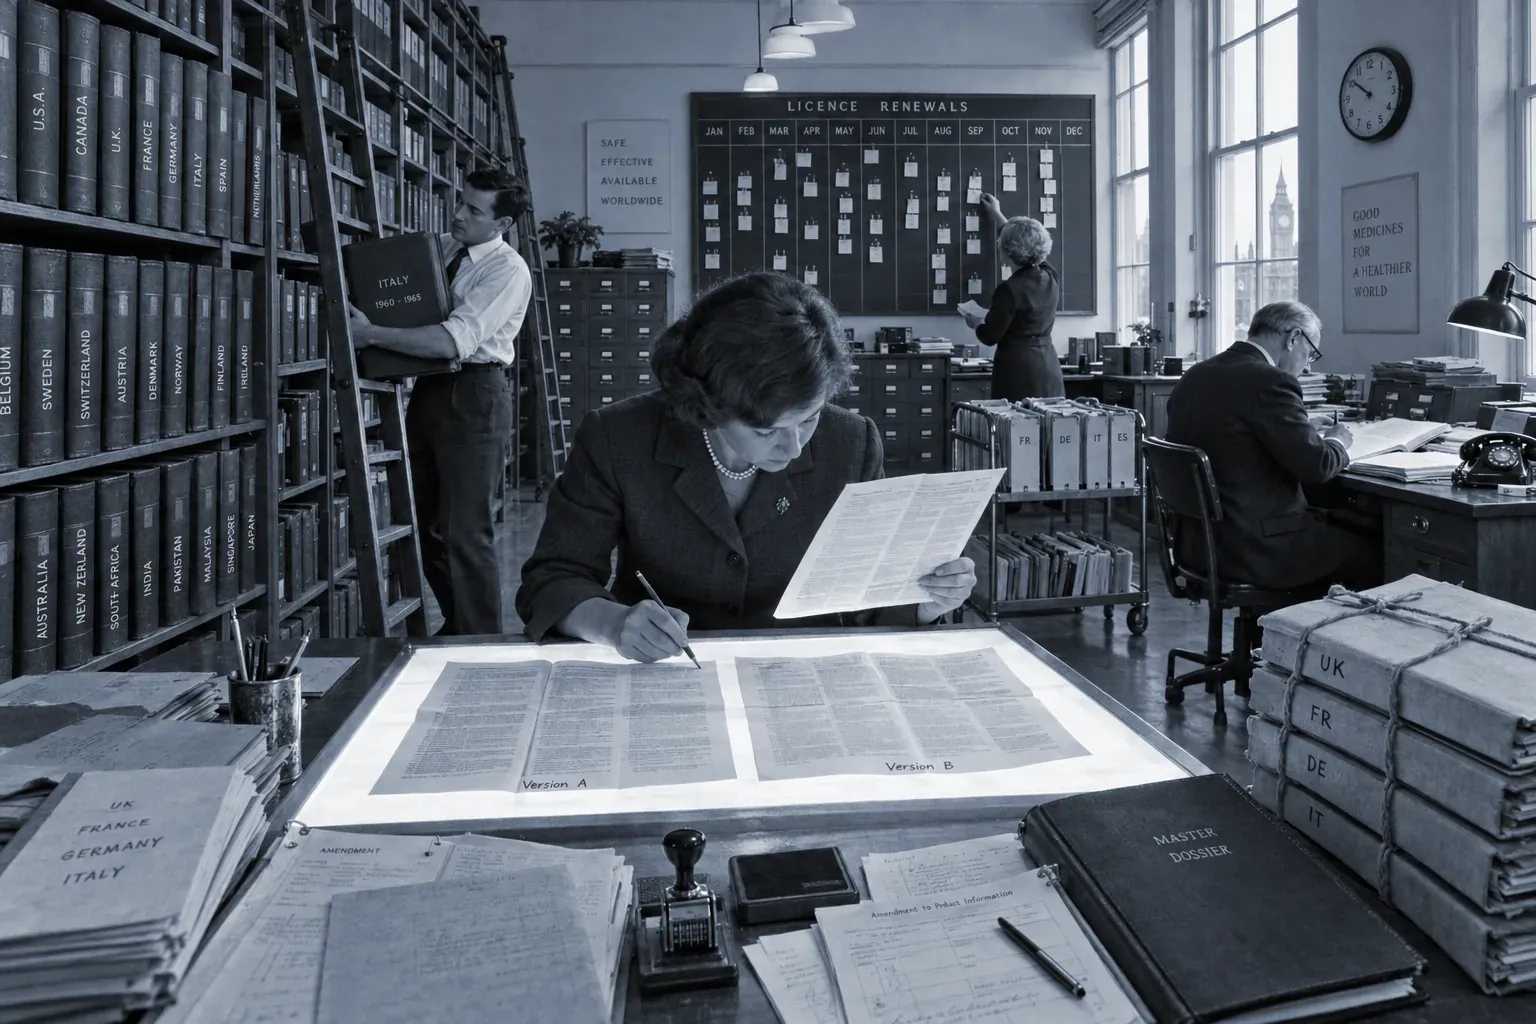
How does lifecycle management keep a marketing authorization current, without variations interrupting supply?
We carry marketing authorizations through lifecycle management: variations under Regulation (EC) No 1234/2008, renewals, PSURs and US supplements under 21 CFR 314.70. How a change is classified decides whether you may ship at once or must wait for agency approval.
Overview
Why do changes to an existing marketing authorization become expensive?
Variations type IA/IAIN/IB/II and extensions under Regulation (EC) No 1234/2008 · renewal under Art. 24 of Directive 2001/83/EC · PSUR under Art. 107b et seq. · PAS, CBE-30 and CBE-0 under 21 CFR 314.70 · changes under EU 2017/745 and EU 2017/746
Last updated: September 16, 2026
The lifecycle is the longest phase of a marketing authorization and the worst resourced. The approval project has a team, the maintenance that follows rarely does. Four places drive the cost.
- Misclassified changes: whether a change counts as type IA, IAIN, IB or II under Regulation (EC) No 1234/2008 determines the filing route and the waiting time. A change classified too low is rejected, and the affected batch stands still until the matter is settled.
- Changes filed one by one instead of grouped: Regulation (EC) No 1234/2008 provides for grouping and work sharing. Filing the same change separately in every country means paying fees and review time repeatedly for one and the same matter.
- Deadlines without an owner: renewal under Article 24 of Directive 2001/83/EC, PSUR dates from the EURD list and the annual report under 21 CFR 314.81(b)(2) each run on their own clock. Nobody notices them until one has been missed.
- Product changes assessed too late in regulatory terms: under EU 2017/745 a change to a product or to the quality management system can trigger notification of the notified body; for devices under the transitional provision of Article 120, classification as a significant change decides whether that provision still applies.
Services
How we support you
Classification and change strategy
Assessment of every planned change against Regulation (EC) No 1234/2008 and Annex I: type IA, IAIN, IB, II or extension, with a reasoned classification, filing route and expected procedural timeline.
US supplements and annual reports
Classification under 21 CFR 314.70 and 601.12 into prior approval supplement, CBE-30 and CBE-0 with the distribution each permits, plus the annual report under 21 CFR 314.81(b)(2).
Renewals and PSURs
Renewal under Article 24 of Directive 2001/83/EC, PSURs under Articles 107b et seq. in the PBRER format per ICH E2C(R2), and scheduling along the EURD list.
Ongoing lifecycle maintenance as a standing service
Taking over the recurring work across a portfolio: change notifications, renewals, reporting obligations and agency questions, with a defined response time rather than a project end date.
How we work together
What it comes down to
A marketing authorization is not a state, it is an inventory that has to be maintained. The effort does not sit where people expect it: not in the dossier but in the classification. Whether a change counts as type IA, IB or II under Regulation (EC) No 1234/2008 decides whether you may ship immediately, wait thirty days, or wait for approval, and the same question arises again, in a different form, in the US as CBE-0, CBE-30 or a prior approval supplement under 21 CFR 314.70.
The second lever is grouping. Sending the same change through twenty countries individually costs twenty times the fee and the review; Articles 7 and 20 of the regulation expressly provide for grouping and work sharing, and for larger portfolios the difference between the two routes is the single biggest item in the annual lifecycle budget.
The third is the deadline without an owner. Renewal under Article 24 of Directive 2001/83/EC, PSUR data lock points from the EURD list, the annual report under 21 CFR 314.81(b)(2): these clocks run on regardless of any project and go unnoticed until one is missed. For medical devices, EU 2017/745 adds that a technical change can trigger something in regulatory terms that development never sees, up to the question of whether the Article 120 transitional provision still holds.
Our approach
Our approach
Step
Result
Portfolio inventory
List of all marketing authorizations and certificates with procedure type, countries, deadlines and open commitments.
Classification of the planned change
Reasoned category under Regulation (EC) No 1234/2008 or 21 CFR 314.70, with filing route and the consequence for distribution.
Grouping and sequencing
Plan for which changes run as a group or under work sharing and in what order, so that dependent changes do not block one another.
Dossier and filing
Variation or supplement filed in the required format, with the affected modules and the rationale for the classification.
Agency questions
Questions answered within the procedural deadlines, with the documented effect on the authorization text and product information.
Updating the records
Updated product information, updated dossier and a refreshed deadline list for renewal and reporting obligations.
Common pitfalls
Where projects commonly fail
A change is classified too low in order to ship sooner.
A change notified as type IA that is a type II change under Annex I of Regulation (EC) No 1234/2008 will be rejected; the goods then stand still longer, not shorter.
Grouping and work sharing go unused.
The same change enters the procedure separately for every country and every authorization, even though Articles 7 and 20 of Regulation (EC) No 1234/2008 expressly provide for bundling.
The renewal is treated as a formality.
Article 24 of Directive 2001/83/EC requires a consolidated version of the dossier; anyone who has filed changes for years without consolidating them is building up work for the renewal date.
PSUR dates are tracked from an internal calendar instead of the EURD list.
The list under Article 107c(7) of Directive 2001/83/EC sets the data lock point and cycle bindingly, and it changes.
For medical devices the change is assessed technically but not in regulatory terms.
Under EU 2017/745 the same design change can trigger notification of the notified body; for devices under Article 120 it decides whether the transitional provision still holds.
Regulatory Affairs
Do any of these pitfalls apply to you?
In a first call we assess your situation and say what needs clarifying first in your case. Without obligation, reply usually within one working day.
FAQ
Frequently asked questions
Sources
- Regulation (EC) No 1234/2008 - primary text including Annex I (variations)
- European Commission guidelines on the details of the various categories of variations (classification guideline)
- Directive 2001/83/EC, Articles 23, 24 and 107b to 107e
- Regulation (EC) No 726/2004 - primary text
- Commission Implementing Regulation (EU) No 520/2012
- ICH E2C(R2): Periodic Benefit-Risk Evaluation Report
- US FDA - 21 CFR 314.70, 314.81, 601.12
- FDA Guidance: Changes to an Approved NDA or ANDA
- Regulation (EU) 2017/745 (MDR), Articles 10, 83 to 86, 120
- Regulation (EU) 2017/746 (IVDR), Article 81
Image notice: The images on this page were generated using artificial intelligence. They do not depict real people, places, or events.
Life Science Journal
Regulatory updates, straight to your inbox.
New requirements, authority decisions and practice notes. Once a month, unsubscribe any time.
Projects from this industry
What we have worked on in this industry
Latest insights
All insights →Regulations & standards considered
- Regulation (EC) No 1234/2008 (examination of variations to marketing authorizations)
- Directive 2001/83/EC, Articles 23 and 24 (obligation to keep current, renewal)
- Directive 2001/83/EC, Articles 107b to 107e (PSUR and PSUSA procedure)
- Regulation (EC) No 726/2004 (centralized procedure, changes to centrally authorized medicines)
- Commission Implementing Regulation (EU) No 520/2012 (pharmacovigilance, PSUR format and content)
- ICH E2C(R2) (Periodic Benefit-Risk Evaluation Report, PBRER)
- 21 CFR 314.70 (Supplements and other changes to an approved NDA: PAS, CBE-30, CBE-0)
- 21 CFR 601.12 (Changes to an approved BLA)
- 21 CFR 314.81(b)(2) (Annual Report)
- EU 2017/745 (MDR), Articles 10, 83 to 86 and 120 (changes, PMS and PSUR)
- EU 2017/746 (IVDR), Article 81 (PSUR for IVDs)
Related topics
Regulatory Affairs →
The authorization itself, before its maintenance begins
Health Authority Interaction →
Questions and procedures with the EMA, FDA and national authorities
Labeling & IFU →
Product information and labeling that every change has to follow
International Market Access →
The same change across several countries and procedures
eCTD Publishing & Regulatory Operations →
The technical form in which every variation reaches the authority
Have a concrete project?
Briefly outline your situation. We'll respond with an initial assessment, usually within one business day.
Prefer direct? +49 89 4161170-0
info@theentourage.de
- Reply usually within one working day
- 4 offices: DE · CH · IT · US
- 100% life sciences